Resource center
What is a medical device? US and EU definitions
What is a medical device depends on the law of the market concerned. This guide sets out the US and EU definitions, the risk classes, and where rapid tests sit.
- Guide
- US and EU sources cited
- Reviewed October 2026
What is a medical device in US law: section 201(h)
The answer to “what is a medical device” is a legal one. In the United States, section 201(h) of the Federal Food, Drug, and Cosmetic Act defines a device as an instrument, apparatus, implement, machine, contrivance, implant, in vitro reagent, or other similar or related article, including a component part or accessory.
Such an article qualifies as a device when it is intended for use in the diagnosis of disease or other conditions, or in the cure, mitigation, treatment or prevention of disease, or to affect the structure or any function of the body. It must not achieve its primary intended purposes through chemical action within or on the body, or depend on being metabolized.
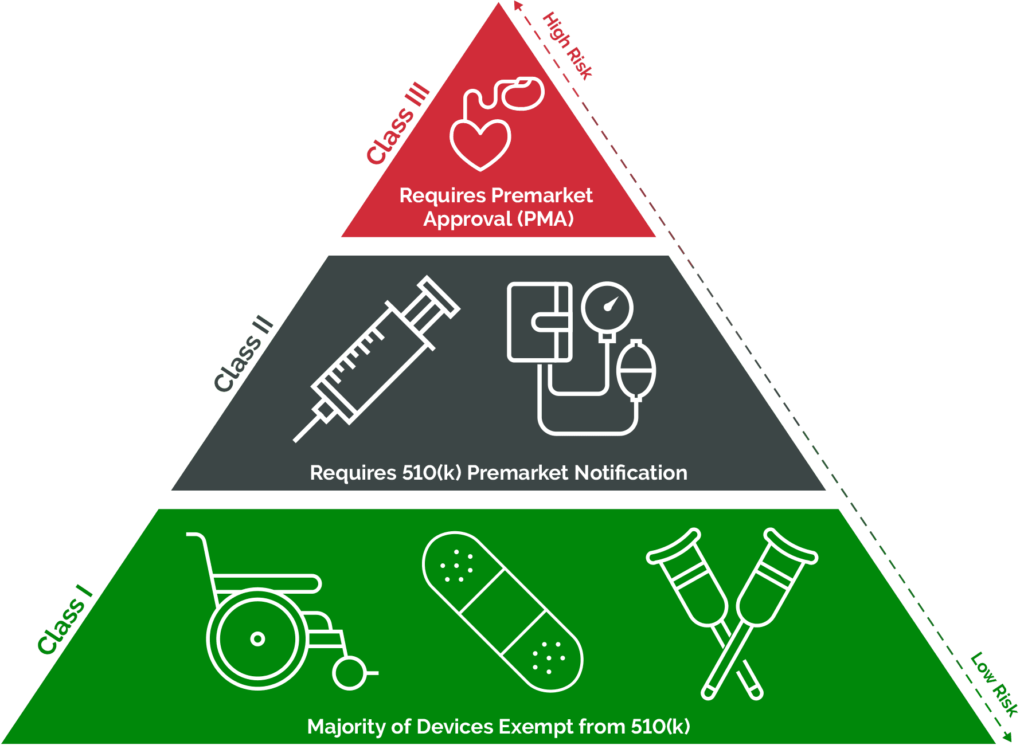
The U.S. Food and Drug Administration (FDA) assigns device types to Class I, II or III. Classification is risk based: Class I includes devices with the lowest risk and Class III those with the greatest.
The EU definition: MDR 2017/745
In the European Union, Regulation (EU) 2017/745 (the MDR) defines a medical device as any instrument, apparatus, appliance, software, implant, reagent, material or other article intended by the manufacturer to be used for human beings for specific medical purposes, such as the diagnosis, prevention, monitoring, treatment or alleviation of disease. It must not achieve its principal intended action by pharmacological, immunological or metabolic means.
Devices fall into four classes: I, IIa, IIb and III. The MDR became applicable on 26 May 2021.
In vitro diagnostics in the EU: IVDR 2017/746 and notified bodies
In vitro diagnostic medical devices have their own regulation, Regulation (EU) 2017/746 (the IVDR); the MDR does not apply to them. The IVDR covers reagents, kits, instruments, software and related articles intended by the manufacturer to be used in vitro for the examination of specimens derived from the human body. It sorts devices into classes A to D, with risk increasing from A to D, and became applicable on 26 May 2022.
Conformity is assessed by notified bodies, organizations designated by EU Member States to assess products before they are placed on the market. Class A devices are self-certified by their manufacturers unless sold sterile, while classes B, C and D require a notified body. Devices in conformity bear the CE mark. This assessment is the task of notified bodies, not of the European Medicines Agency.
US and EU systems at a glance
US and EU compared
| System | Legal basis | Risk classes | Pre-market review |
|---|---|---|---|
| United States | FD&C Act, section 201(h) | Class I, II, III | FDA: 510(k) for non-exempt Class I or II devices; premarket approval for Class III |
| EU medical devices | Regulation (EU) 2017/745 | Class I, IIa, IIb, III | Notified body, generally not for Class I |
| EU in vitro diagnostics | Regulation (EU) 2017/746 | Class A, B, C, D | Notified body for classes B, C and D |
Simplified summary of the sources listed under References. Not regulatory advice.
Where rapid tests sit
Rapid tests examine a specimen outside the body, so they are in vitro diagnostic devices in both systems. FDA states that in vitro diagnostic products are devices as defined in section 201(h) and classifies them into Class I, II or III. In the EU they fall under the IVDR, which defines devices for self-testing by lay persons separately from devices for near-patient testing by health professionals.
Regulatory status is decided product by product and market by market. First Diagnostic™ Corporation brands and distributes lateral flow tests made by established IVD manufacturers, for professional in vitro diagnostic use. See the rapid tests catalog, such as the hepatitis and HIV rapid tests, and our quality and regulatory information. The retail pharmacy channel guide covers self-tests. For a product’s status in your market, contact us.
Frequently asked questions
Is a rapid test a medical device?
Yes. It is an in vitro diagnostic device in both the US and the EU systems.
Does the European Medicines Agency approve medical devices?
No. In the EU, conformity is assessed by notified bodies, or declared by the manufacturer for the lowest-risk classes.
Does a CE mark allow sale in every country?
No. CE marking relates to EU requirements. Countries outside the EU set their own rules, and many require a separate national registration.
Keep reading
- Quality and regulatory information — How First Diagnostic handles documentation.
- What are rapid diagnostic tests (RDTs)? — Formats, analytes and evaluation points.
- The retail pharmacy channel for rapid tests — Professional-use and self-test products.
- What is a manufacturer’s representative? — Sales roles compared with regulatory roles.
References
- U.S. Food and Drug Administration. How to Determine if Your Product is a Medical Device (content current as of 29 September 2022). fda.gov
- U.S. Food and Drug Administration. Classify Your Medical Device. fda.gov
- U.S. Food and Drug Administration. Overview of IVD Regulation (content current as of 20 December 2024). fda.gov
- Regulation (EU) 2017/745 of the European Parliament and of the Council of 5 April 2017 on medical devices, Articles 1 and 2 (consolidated text of 24 April 2020). EUR-Lex. eur-lex.europa.eu
- Regulation (EU) 2017/746 of the European Parliament and of the Council of 5 April 2017 on in vitro diagnostic medical devices, Article 2. EUR-Lex. eur-lex.europa.eu
- European Commission. Notified bodies for medical devices. health.ec.europa.eu
- European Commission. Factsheet for healthcare professionals and health institutions (18 December 2024). health.ec.europa.eu
- European Commission. Factsheet for manufacturers of in vitro diagnostic medical devices (1 August 2020). health.ec.europa.eu
Source: First Diagnostic™ editorial team · Last reviewed: October 2026 · First Diagnostic™ editorial team
Regulatory status: Products shown on this website are for professional in vitro diagnostic use. They are not cleared, approved or authorized by the U.S. Food and Drug Administration and are not offered for sale in the United States. Product availability and regulatory status vary by country; contact us for the status in your market.